A vast landscape of ‘undruggable’ cancer targets remains beyond the reach of conventional therapeutic agents. Recent advances in artificial intelligence (AI), however, are challenging this paradigm. Synthesizing insights from a Cancer Moonshot workshop, we argue that systemically addressing the undruggable target space with AI requires a new conceptual framework. We highlight the failure of current target taxonomies and the need for benchmarking datasets, and re-evaluate clinical validation for novel AI-driven modalities.
J Clin Microbiol. 2025 Aug 19:e0058525. doi: 10.1128/jcm.00585-25. Online ahead of print.
ABSTRACT
Human papillomavirus (HPV) is comprised of >200 genotypes and has an ~8 kb, circular, double-stranded DNA genome. Transmission of HPV occurs through skin-to-skin contact and infection of squamous epithelial cells of cutaneous and mucosal surfaces. HPV genotypes are categorized as low- or high-risk (hrHPV) based on oncogenic potential. There are approximately 14 types of hrHPV that can cause several types of cancer, including HPV-associated oropharyngeal squamous cell carcinoma (HPV(+)OPSCC). Detection of HPV(+)OPSCC is traditionally accomplished using p16 immunohistochemistry (IHC) and HPV-specific testing, either DNA or RNA in situ hybridization (ISH) staining or DNA-based PCR of suspected tumor biopsy tissue. More recently, platelet-poor plasma (PPP) samples from patients with HPV(+)OPSCC have proven useful for detection and quantitation of fragments of HPV circulating tumor DNA (ctDNA). ctDNA has been shown to be useful in determining treatment response and monitoring for disease recurrence. In this study, a novel droplet digital PCR assay (ddPCR) was developed and validated for the detection and quantitation of ctDNA from 5 hrHPV genotypes in PPP. Analytical sensitivity ranged from 7.71 to 19.45 fragments of HPV ctDNA per milliliter of PPP across five hrHPV genotypes. In patients with confirmed primary or recurrent HPV(+)OPSCC or HPV(-)OPSCC, testing of corresponding PPP samples (n = 32) by ddPCR demonstrated 90.63% (29/32) overall agreement with p16/HPV-ISH biopsy results. Compared with reference ddPCR assays performed at outside laboratories, our ddPCR assay yielded 90% (9/10) overall agreement. This assay may provide clinicians with a tool for monitoring HPV ctDNA prior to, during, and after treatment of an HPV-associated cancer.
IMPORTANCE: At least 14 genotypes of human papillomavirus (HPV) have been identified to have high oncogenic potential. While molecular diagnostic testing for HPV is widely available for liquid cytologic cervical samples, testing is limited for other sample types, including liquid biopsy samples, such as platelet-poor plasma (PPP). With the rising incidence of HPV-associated oropharyngeal squamous cell carcinoma (HPV(+)OPSCC), laboratory testing is an essential part of patient diagnosis, management, and surveillance. Here, we summarize the development and analytical performance validation of a multiplexed, droplet digital PCR (ddPCR) assay for the detection and quantitation of HPV circulating tumor DNA (ctDNA) in PPP. This assay may provide clinicians with a tool to address minimal residual disease for patients with an HPV-associated cancer.
Expert Rev Anticancer Ther. 2025 Aug 18. doi: 10.1080/14737140.2025.2549538. Online ahead of print.
ABSTRACT
INTRODUCTION: In the era of precision medicine, molecular biomarker testing is increasingly becoming standard of care for Non-Small Cell Lung Cancer (NSCLC) patients. Tissue and liquid biopsy-based Next-Generation Sequencing (NGS) is now highly recommended.
AREAS COVERED: Different NGS platforms emerged as a cost-effective strategy to perform a massive and parallel sequencing performing higher technical sensitivity than old generation technologies in detecting low abundant alterations in challenging diagnostic samples. NGS systems can detect single nucleotide variants (SNV), small insertions and deletions (indels), copy number alterations (CNAs) and structural variants (SVs) or gene fusions across selected druggable genes optimizing clinical administration of NSCLC patients. The diagnostic implementation of the most adequate NGS panel depending on several factors that could impact on the clinical utility of testing assay.
EXPERT OPINION: Promising advanced technologies are emerging as potentially integrative tools in personalized medicine. In this context, multi-omic evaluation including genomic, transcriptomic, fragmentomic and epigenomic signatures are under investigation to significantly modify clinical algorithm of NSCLC patients. On this basis, sequencing strategies may play a pivotal role in the implementation of a new predictive model for cancer diagnosis and prognosis.
Victor Dibia shares insights into multi-agent systems. He explains their definition, demonstrates implementation using the AutoGen framework, and identifies 10 common reasons these systems fail in production. He provides guidance on when a multi-agent approach is appropriate, emphasizing evaluation-driven design and tool-focused implementations.
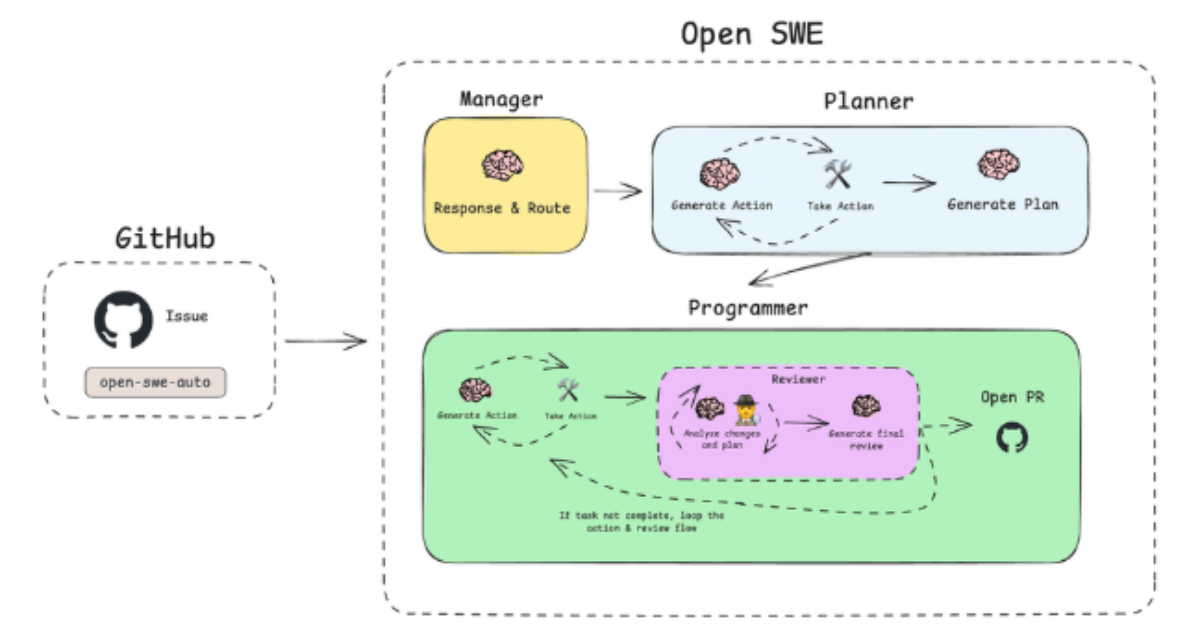
LangChain has released Open SWE, a fully open-source, asynchronous coding agent designed to operate in the cloud and handle complex software development tasks. The company says Open SWE represents a shift away from real-time “copilot” assistants toward more autonomous, long-running agents that integrate directly with a developer’s existing workflows.
A study presents ALPACA, a computational method for inferring clone- and allele-specific copy numbers of individual clones from multi-sample bulk DNA-sequencing data, and demonstrates its use to study metastasis trajectories.
Artificial intelligence models that can discover drugs and write code still fail at puzzles a lay person can master in minutes. This phenomenon sits at the heart of the challenge of artificial general intelligence (AGI). Can today’s AI revolution produce models that rival or surpass human intelligence across all domains? If so, what underlying enablers—whether hardware, software, or the orchestration of both—would be needed to power them?
Dario Amodei, co-founder of Anthropic, predicts some form of “powerful AI” could come as early as 2026, with properties that include Nobel Prize-level domain intelligence; the ability to switch between interfaces like text, audio, and the physical world; and the autonomy to reason toward goals, rather than responding to questions and prompts as they do now. Sam Altman, chief executive of OpenAI, believes AGI-like properties are already “coming into view,” unlocking a societal transformation on par with electricity and the internet. He credits progress to continuous gains in training, data, and compute, along with falling costs, and a socioeconomic value that is “super-exponential.”
Optimism is not confined to founders. Aggregate forecasts give at least a 50% chance of AI systems achieving several AGI milestones by 2028. The chance of unaided machines outperforming humans in every possible task is estimated at 10% by 2027, and 50% by 2047, according to one expert survey. Time horizons shorten with each breakthrough, from 50 years at the time of GPT-3’s launch to five years by the end of 2024. “Large language and reasoning models are transforming nearly every industry,” says Ian Bratt, vice president of machine learning technology and fellow at Arm.
This content was produced by Insights, the custom content arm of MIT Technology Review. It was not written by MIT Technology Review’s editorial staff.
This content was researched, designed, and written entirely by human writers, editors, analysts, and illustrators. This includes the writing of surveys and collection of data for surveys. AI tools that may have been used were limited to secondary production processes that passed thorough human review.
Cell Cycle. 2025 Aug 13:1-22. doi: 10.1080/15384101.2025.2539645. Online ahead of print.
ABSTRACT
Epithelial carcinoma cells require penetration of the basement membrane (BM) to metastasize. The BM is a thin layer of extracellular matrix beneath epithelial and endothelial tissues. It acts as a structural barrier, preventing cancer cells from invading and undergoing endocytosis and exocytosis. Thus, understanding the relationship between the BM and tumor immunity can lead to new strategies for halting cancer progression and metastasis. Gene expression data of 33 cancers were obtained from the Cancer Genome Atlas database. The study analyzed the correlation between BM regulatory genes, copy number variations, immune-related genes, and tumor immune dysfunction rejection (TIDE). Immunohistochemical methods were used to analyze the expression of regulatory genes. And the BM score was calculated using single-sample gene set enrichment analysis. Single-cell transcriptional sequencing determined the activation status of the BM in the tumor microenvironment. The expression of BM-related genes (BMGs) exhibited significant heterogeneity across different cancer types. Most genes were up-regulated in tumor tissues. Major single nucleotide polymorphisms of BMGs included missense mutations, while major copy number variations were heterozygous deletion and heterozygous amplification. Additionally, the expressions of immune checkpoint molecules CD276, NRP1, and C10orf54 showed positive correlations with BMS. Numerous tumors displayed a significant positive correlation between BMS and TIDE scores. We demonstrate that BM regulatory genes undergo alterations specific to different cancer types, which are associated with the expression of immune checkpoints and immune dysfunction. This indicates that BM remodeling plays an active role in modulating immune resistance, rather than being a passive structural alteration.
Biomark Res. 2025 Aug 11;13(1):101. doi: 10.1186/s40364-025-00812-z.
ABSTRACT
BACKGROUND: Early identification of high-risk individuals is crucial for optimizing cancer screening, particularly when considering expensive and invasive methods such as multi-omics technologies and endoscopic procedures. However, developing a robust, practical multi-cancer risk prediction model that integrates diverse, multi-scale data and with proper validation remains a significant challenge.
METHODS: We initialized the FuSion study by recruiting 42,666 participants from Taizhou, China, with a discovery cohort (n = 16,340) and an independent validation cohort (n = 26,308) after exclusion criteria. We integrated multi-scale data from 54 blood-derived biomarkers and 26 epidemiological exposures to develop a risk prediction model for five common cancers, including lung, esophageal, liver, gastric, and colorectal cancer. Employing five supervised machine learning approaches, we used a LASSO-based feature selection strategy to identify the most informative predictors. The model was trained and internally validated in the discovery cohort, externally applied in the validation cohort, and further evaluated through a prospective clinical follow-up to assess cancer events via clinical examinations.
RESULTS: The final model comprising four key biomarkers along with age, sex, and smoking intensity, achieving an AUROC of 0.767 (95% CI: 0.723-0.814) for five-year risk prediction. High-risk individuals (17.19% of the cohort) accounted for 50.42% of incident cancer cases, with a 15.19-fold increased risk compared to the low-risk group. During follow-up of 2,863 high-risk subjects, 9.64% were newly diagnosed with cancer or precancerous lesions. Notably, cancer detection in the high-risk group was 5.02 times higher than in the low-risk group and 1.74 times higher than in the intermediate-risk group. In particular, the incidence of esophageal cancers in the high-risk group was 16.84 times that of the low-risk group.
CONCLUSIONS: This is the first population-based prospective study in a large Chinese cohort that leverage multi-scale data including biomarkers for multi-cancer risk prediction. Our effective risk stratification model not only enhances early cancer detection but also lays the foundation for the targeted application of advanced screening methods, including but not limited to multi-omics technologies and endoscopy. These findings support precision prevention strategies and the optimal allocation of healthcare resources.
J Transl Med. 2025 Aug 7;23(1):877. doi: 10.1186/s12967-025-06725-7.
ABSTRACT
BACKGROUND: Gastric cancer (GC) is characterized by significant intertumoral heterogeneity, which often leads to the development of resistance to platinum-based chemotherapy. Combining platinum drugs with other therapeutic strategies may improve treatment efficacy; however, the mechanisms underlying platinum resistance in GC remain unclear.
METHODS: Key genes related to platinum resistance in GC were selected from the platinum resistance gene database and GC resistance datasets. The Similarity Network Fusion (SNF) algorithm was employed, along with prognosis-related methylation data and somatic mutation data, to classify the molecular subtypes of GC based on GC platinum resistance genes. Gene expression profiles, prognosis, immune cell infiltration, chemotherapy sensitivity, and immunotherapy responsiveness were comprehensively evaluated for each subtype. Localization and functional evaluation were conducted at the single-cell and spatial transcriptomics levels, and predictive models were developed using machine learning techniques. These functional differences in platinum resistance gene models were further explored in GC. Moreover, experimental validation was conducted to elucidate the mechanisms of key genes involved in platinum resistance in GC.
RESULTS: Stomach adenocarcinoma (STAD) patients were classified into three subtypes using the SNF algorithm and multiomics data. Patients with subtype CS2 exhibited a significantly poorer prognosis than those with subtypes CS1 and CS3 (p < 0.05). Subtype CS1 was characterized as immune-deprived, CS2 as stroma-enriched, and CS3 as immune-enriched. Patients with subtype CS2 also exhibited the most adverse therapeutic responses to docetaxel, cisplatin, and gemcitabine. Single-cell analysis revealed high enrichment of M1 module cells with elevated expression of resistance genes, including the transcription factor KLF9. Spatial transcriptomic analysis further confirmed the independent spatial distribution of malignant cells with high expression of drug resistance genes (DRGs). Predictive models based on machine learning demonstrated excellent prognostic performance. Patients in the high DRG group also exhibited poorer responses to immunotherapy. Cellular experiments revealed that KLF9 overexpression significantly inhibited the proliferation of AGS cells (p < 0.05), reduced their resistance to platinum-based drugs, and markedly decreased the levels of inflammatory cytokines in them.
CONCLUSION: KLF9 was identified as a promising therapeutic target for overcoming platinum resistance in GC, warranting further investigation into its role and potential clinical applications.
AIM: This study aims to present a comprehensive international analysis of the existing techniques used in liquid biopsies and their use in isolating tumor markers to detect, predict, and monitor the results of cancer treatment.
MATERIALS AND METHODS: We conducted a narrative review using a scoping review model based on three databases, including PubMed/Medline, Scopus, and Cochrane. The search criteria included all articles on liquid biopsy of the last five years (June 30th, 2023-Oct 30, 2024) ((liquid Biopsy) AND (("2023/06/30"[Date - Publication]: "2024/10/30"[Date - Publication]))). We also approached gray literature on this topic. We focused on review articles as an eligibility criterion for this narrative review, but we also carried out a United States registered clinical trials review targeting immunotherapy and liquid biopsy with the limitation "recruiting" and/or "not yet recruiting" (updated on March 31, 2025).
RESULTS: We screened 2645 articles from PubMed/Medline, Scopus, and Cochrane and 45 articles from the gray literature. We retrieved the full text for 325 articles. Liquid biopsies involve the extraction of tumor-derived components such as circulating tumor cells, circulating tumor DNA, and tumor extracellular vesicles from the bodily fluids of cancer patients. We found 25 United States registered governmental clinical trials targeting immunotherapy and liquid biopsy, of which 20 trials are recruiting and five trials are not yet recruiting.
DISCUSSION: Developments in medicine have led to a more comprehensive understanding of tumor features, including tumor load, tumor staging, heterogeneity, gene mutations, and clonal evolution. The utilization of liquid biopsies from cancer patients has provided novel opportunities for detection and ongoing monitoring, precision medicine-based therapy, and identification of markers for therapeutic resistance.
Commun Biol. 2025 Aug 6;8(1):1160. doi: 10.1038/s42003-025-08533-7.
ABSTRACT
The rapid advancement of single-cell technologies has led to the development of various analysis methods, each with trade-offs between predictive power and interpretability particularly for multimodal data integration. Complex machine learning models achieve high accuracy, but they often lack transparency, while simpler models are more interpretable but less effective for prediction. In this manuscript, we introduce an innovative method for single-cell analysis using Multiple Kernel Learning (scMKL), that merges the predictive capabilities of complex models with the interpretability of linear approaches, aimed at providing actionable insights from single-cell multiomics data. scMKL excels at classifying healthy and cancerous cell populations across multiple cancer types, utilizing data from single-cell RNA sequencing, ATAC sequencing, and 10x Multiome. It outperforms existing methods while delivering interpretable results that identify key transcriptomic and epigenetic features, as well as multimodal pathways- that existing methods have failed to achieve, in breast, lymphatic, prostate, and lung cancers. Leveraging insights from one dataset to inform analysis in a new dataset, scMKL uncovers biological pathways that distinguish treatment responses in breast cancer, low-grade from high-grade prostate tumors, and subtypes in lung cancer, thereby enhancing our understanding of cancer biology and tumor progression.
Int J Med Sci. 2025 Jul 10;22(13):3268-3276. doi: 10.7150/ijms.117603. eCollection 2025.
ABSTRACT
Hepatocellular carcinoma (HCC) is a genetically heterogeneous malignancy in which single-gene mutations serve as critical drivers of tumor initiation, progression, and therapeutic resistance. Advances in high-throughput genomics and liquid biopsy technologies have highlighted the clinical utility of mutations in genes such as TP53, CTNNB1, and TERT as diagnostic, prognostic, and predictive biomarkers. These mutations disrupt key oncogenic pathways, modulate the tumor immune microenvironment, and contribute to intratumoral heterogeneity, complicating disease management. Mutation-guided precision medicine, including telomerase inhibitors, Wnt/β-catenin pathway modulators, and immune checkpoint blockade, offers promising avenues for individualized treatment in HCC. However, challenges persist in translating these findings into clinical practice due to mutation complexity, resistance mechanisms, and limitations in biomarker standardization. Emerging strategies such as multi-omics integration, artificial intelligence, and gene editing technologies hold potential to overcome these barriers and facilitate the development of personalized therapeutic regimens. This review summarizes the molecular mechanisms, clinical applications, and translational challenges of single-gene mutations in HCC, with the aim of guiding future research and precision oncology.
Nat Commun. 2025 Aug 5;16(1):7178. doi: 10.1038/s41467-025-62593-1.
ABSTRACT
Spatial transcriptomics links gene expression with tissue morphology, however, current tools often prioritize genomic analysis, lacking integrated image interpretation. To address this, we present Thor, a comprehensive platform for cell-level analysis of spatial transcriptomics and histological images. Thor employs an anti-shrinking Markov diffusion method to infer single-cell spatial transcriptome from spot-level data, effectively combining gene expression and cell morphology. The platform includes 10 modular tools for genomic and image-based analysis, and is paired with Mjolnir, a web-based interface for interactive exploration of gigapixel images. Thor is validated on simulated data and multiple spatial platforms (ISH, MERFISH, Xenium, Stereo-seq). Thor characterizes regenerative signatures in heart failure, screens breast cancer hallmarks, resolves fine layers in mouse olfactory bulb, and annotates fibrotic heart tissue. In high-resolution Visium HD data, it enhances spatial gene patterns aligned with histology. By bridging transcriptomic and histological analysis, Thor enables holistic tissue interpretation in spatial biology.
A study reports whole-genome sequences for 490,640 participants from the UK Biobank and combines these data with phenotypic data to provide new insights into the relationship between human variation and sequence variation.
Future Oncol. 2025 Aug 5:1-19. doi: 10.1080/14796694.2025.2542051. Online ahead of print.
ABSTRACT
Lung Cancer (LC) remains the leading cause of cancer-related mortality. While Tissue Biopsy (TB) remains the gold standard for molecular profiling, its invasiveness and inability to provide real-time monitoring have led to the adoption of Liquid Biopsy (LB) as a minimally invasive alternative. By analyzing different circulating analytes such as cell-free DNA (cfDNA), circulating tumor DNA (ctDNA), Circulating Tumor Cells (CTCs), Extracellular Vesicles (EVs), and Tumor-Educated Platelets (TEPs), LB offers a dynamic approach to assessing tumor heterogeneity, Minimal Residual Disease (MRD), and treatment resistance. Recent clinical trials have underscored their role in guiding therapy decisions and monitoring treatment response. In early-stage disease, several Randomized Clinical Trials (RCTs) have shown that ctDNA clearance predicts survival benefits in patients receiving neoadjuvant or perioperative Immune Checkpoint Inhibitors (ICIs). Additionally, adjuvant RCTs have confirmed the ctDNA prognostic role in post-surgical relapse risk assessment. Despite its transformative potential, challenges such as assay standardization, sensitivity limitations in early-stage disease, and regulatory barriers remain. As ongoing research continues to validate its clinical utility, LB is poised to become an indispensable tool in the precision management of LC.
Eur Urol Focus. 2025 Aug 1:S2405-4569(25)00178-6. doi: 10.1016/j.euf.2025.06.009. Online ahead of print.
ABSTRACT
BACKGROUND AND OBJECTIVE: Urinary tumor DNA (utDNA) has emerged as a promising biomarker in the care, diagnosis, early detection, recurrence monitoring, and prognosis of bladder cancer (BCa). Its noninvasive nature, ease of access, and cost effectiveness make it an attractive option for both patients and health care providers. This review describes the current state of utDNA as a marker of BCa.
METHODS: Articles published between 2015 and 2025 on current utDNA-based techniques in BCa were identified and analyzed for relevance and insight into utDNA research and usage.
KEY FINDINGS AND LIMITATIONS: Recent investigations underscore the noninvasiveness and superior tumor detection capabilities of utDNA, particularly in the detection of minimal residual disease. Moreover, utDNA provides actionable information, such as tumor grade and staging information, to support precise treatment decisions, including targeted immunotherapy regimens and bladder preservation strategies. Although utDNA has shown promising results in small studies, larger studies must be performed before it can be considered as a standard procedure in clinical practice.
CONCLUSIONS AND CLINICAL IMPLICATIONS: Urinary tumor DNA has demonstrated great potential to improve on most, if not all, stages of detection, treatment, and monitoring of BCa. By preserving the low cost and noninvasiveness of urine cytology, and by replacing its suboptimal accuracy with a precision rivaling and often exceeding cystoscopy and circulating tumor DNA-based methods, utDNA offers patients a more comfortable, repeatable, and accurate way of detecting BCa. With increased sensitivity and accuracy, everything from low-grade tumors to the earliest signs of recurrence can be detected more effectively, optimizing patient treatment courses and improving outcomes.

 Login