Normal view
-
Journal of Medical Internet Research
-
Digital Health Technology Infrastructure Challenges to Support Health Equity in the United States: Scoping Review
Background: Even though Digital Health Technology (DHT) is widely utilized in the United States (U.S.) at both hospital provider and individual levels, it is beset with several challenges that have contributed to inequities in the health service delivery. Previous studies have shown that health inequities observed may be amplified many by DHT requirements. Objective: The objectives of this scoping review are aimed at synthesizing information on DHT inequities by exploring evidence that describes
-
InfoQ

-
Hugging Face Releases FinePDFs: a 3-Trillion-Token Dataset Built from PDFs
Hugging Face has unveiled FinePDFs, the largest publicly available corpus built entirely from PDFs. The dataset spans 475 million documents in 1,733 languages, totaling roughly 3 trillion tokens. At 3.65 terabytes in size, FinePDFs introduces a new dimension to open training datasets by tapping into a resource long considered too complex and expensive to process. By Robert Krzaczyński
Hugging Face Releases FinePDFs: a 3-Trillion-Token Dataset Built from PDFs
Hugging Face has unveiled FinePDFs, the largest publicly available corpus built entirely from PDFs. The dataset spans 475 million documents in 1,733 languages, totaling roughly 3 trillion tokens. At 3.65 terabytes in size, FinePDFs introduces a new dimension to open training datasets by tapping into a resource long considered too complex and expensive to process.
By Robert Krzaczyński-
MRD
-
Prognostic Value of Circulating Tumor DNA in HR+/HER2- Stage I-III Breast Cancer: A Systematic Review
Cancers (Basel). 2025 Aug 29;17(17):2831. doi: 10.3390/cancers17172831.ABSTRACTBackground: Hormone receptor-positive (HR+), HER2-negative breast cancer accounts for the majority of breast cancer diagnoses. While outcomes have improved with neoadjuvant and adjuvant therapies, the risk of late recurrence persists, and there remains a critical need for reliable biomarkers to guide prognosis and post-treatment surveillance. Circulating tumor DNA (ctDNA), detectable via liquid biopsy, has emerged as
Prognostic Value of Circulating Tumor DNA in HR+/HER2- Stage I-III Breast Cancer: A Systematic Review
Cancers (Basel). 2025 Aug 29;17(17):2831. doi: 10.3390/cancers17172831.
ABSTRACT
Background: Hormone receptor-positive (HR+), HER2-negative breast cancer accounts for the majority of breast cancer diagnoses. While outcomes have improved with neoadjuvant and adjuvant therapies, the risk of late recurrence persists, and there remains a critical need for reliable biomarkers to guide prognosis and post-treatment surveillance. Circulating tumor DNA (ctDNA), detectable via liquid biopsy, has emerged as a promising tool for monitoring minimal residual disease and predicting survival outcomes. This systematic review evaluates the association between ctDNA detection during neoadjuvant or adjuvant treatment and survival outcomes in early-stage HR+/HER2- breast cancer. Methods: This systematic review was conducted in accordance with PRISMA guidelines. A comprehensive literature search of Ovid MEDLINE and Embase was conducted to identify studies published through 3 May 2024 that evaluated ctDNA as a prognostic biomarker in stage I-III HR+/HER2- breast cancer. We included studies reporting recurrence-free survival, invasive disease-free survival, or overall survival and excluded non-original studies, conference abstracts, and non-English articles. Data extraction and qualitative synthesis were performed, and the risk of bias was qualitatively assessed across studies. No review protocol was registered. Results: Eleven studies comprising 1644 patients met the inclusion criteria. In the neoadjuvant setting, ctDNA positivity prior to treatment initiation was associated with inferior survival outcomes. In the adjuvant setting, detection of ctDNA during or after treatment was consistently linked to poorer recurrence-free and invasive disease-free survival. Across studies, ctDNA detection was a significant negative prognostic marker. Conclusions: This systematic review supports the prognostic value of ctDNA in HR+/HER2- early-stage breast cancer. Limitations include small sample sizes, observational study designs, and heterogeneity in ctDNA assays. Standardization of ctDNA testing methods and further prospective trials are needed to validate its clinical utility and explore its potential role in guiding therapeutic interventions.
PMID:40940926 | PMC:PMC12427406 | DOI:10.3390/cancers17172831
-
Journal of Medical Internet Research
-
Interventions Based on Biofeedback Systems to Improve Workers’ Psychological Well-Being, Mental Health, and Safety: Systematic Literature Review
Background: In modern, high-speed work settings, the significance of mental health disorders is increasingly acknowledged as a pressing health issue, with potential adverse consequences for organizations, including reduced productivity and increased absenteeism. Over the past few years, various mental health management solutions, such as biofeedback applications, have surfaced as promising avenues to improve employees’ mental well-being. However, most studies on these interventions have been con
Interventions Based on Biofeedback Systems to Improve Workers’ Psychological Well-Being, Mental Health, and Safety: Systematic Literature Review
-
Nature - Issue - nature.com science feeds
-
Fluctuating DNA methylation tracks cancer evolution at clinical scale
Nature, Published online: 10 September 2025; doi:10.1038/s41586-025-09374-4Cancer evolutionary dynamics are quantitatively inferred using a method, EVOFLUx, applied to fluctuating DNA methylation.
Fluctuating DNA methylation tracks cancer evolution at clinical scale
Nature, Published online: 10 September 2025; doi:10.1038/s41586-025-09374-4
Cancer evolutionary dynamics are quantitatively inferred using a method, EVOFLUx, applied to fluctuating DNA methylation.-
InfoQ

-
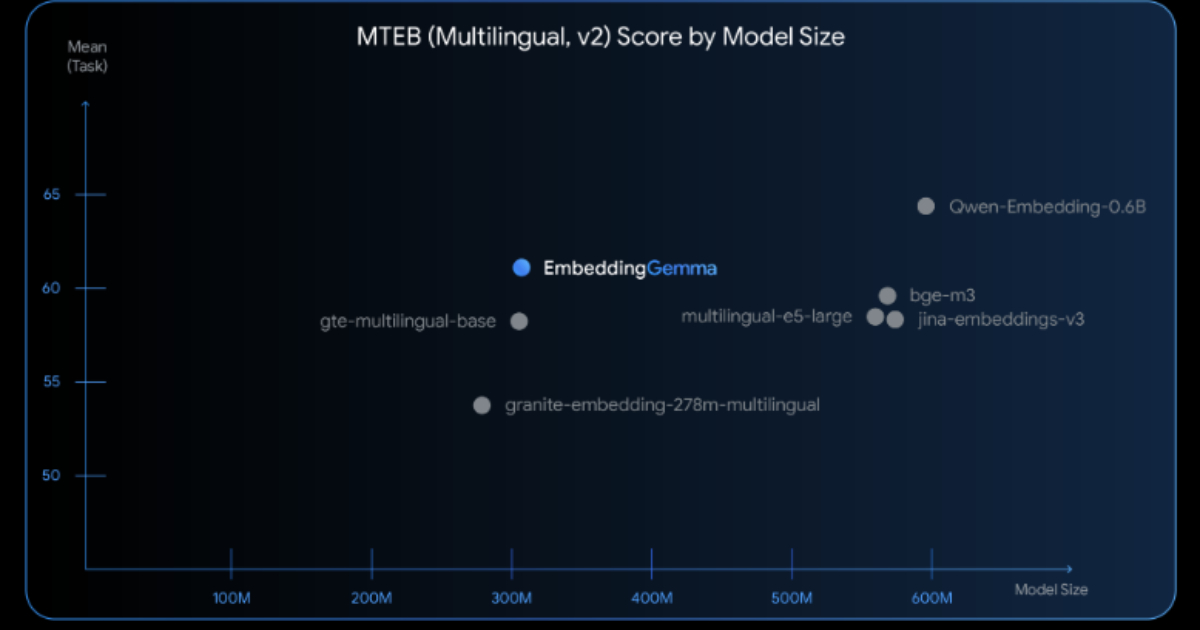
Google DeepMind Launches EmbeddingGemma, an Open Model for On-Device Embeddings
Google DeepMind has introduced EmbeddingGemma, a 308M parameter open embedding model designed to run efficiently on-device. The model aims to make applications like retrieval-augmented generation (RAG), semantic search, and text classification accessible without the need for a server or internet connection. By Robert Krzaczyński
Google DeepMind Launches EmbeddingGemma, an Open Model for On-Device Embeddings
Google DeepMind has introduced EmbeddingGemma, a 308M parameter open embedding model designed to run efficiently on-device. The model aims to make applications like retrieval-augmented generation (RAG), semantic search, and text classification accessible without the need for a server or internet connection.
By Robert Krzaczyński-
(Multiomics OR Omics) AND (Pancreatic)
-
Single-cell multiome and spatial profiling reveals pancreas cell type-specific gene regulatory programs of type 1 diabetes progression
Sci Adv. 2025 Sep 12;11(37):eady0080. doi: 10.1126/sciadv.ady0080. Epub 2025 Sep 10.ABSTRACTCell type-specific regulatory programs that drive type 1 diabetes (T1D) in the pancreas are poorly understood. Here, we performed single-nucleus multiomics and spatial transcriptomics in up to 32 nondiabetic (ND), autoantibody-positive (AAB+), and T1D pancreas donors. Genomic profiles from 853,005 cells mapped to 12 pancreatic cell types, including multiple exocrine subtypes. β, Acinar, and other cell typ
Single-cell multiome and spatial profiling reveals pancreas cell type-specific gene regulatory programs of type 1 diabetes progression
Sci Adv. 2025 Sep 12;11(37):eady0080. doi: 10.1126/sciadv.ady0080. Epub 2025 Sep 10.
ABSTRACT
Cell type-specific regulatory programs that drive type 1 diabetes (T1D) in the pancreas are poorly understood. Here, we performed single-nucleus multiomics and spatial transcriptomics in up to 32 nondiabetic (ND), autoantibody-positive (AAB+), and T1D pancreas donors. Genomic profiles from 853,005 cells mapped to 12 pancreatic cell types, including multiple exocrine subtypes. β, Acinar, and other cell types, and related cellular niches, had altered abundance and gene activity in T1D progression, including distinct pathways altered in AAB+ compared to T1D. We identified epigenomic drivers of gene activity in T1D and AAB+ which, combined with genetic association, revealed causal pathways of T1D risk including antigen presentation in β cells. Last, single-cell and spatial profiles together revealed widespread changes in cell-cell signaling in T1D including signals affecting β cell regulation. Overall, these results revealed drivers of T1D in the pancreas, which form the basis for therapeutic targets for disease prevention.
PMID:40929272 | PMC:PMC12422192 | DOI:10.1126/sciadv.ady0080
-
Nature Medicine
-
The WHO global landscape of cancer clinical trials
Nature Medicine, Published online: 09 September 2025; doi:10.1038/s41591-025-03926-xThis Review of the WHO’s International Clinical Trials Registry Platform presents a snapshot of the global cancer trial landscape and provides critical empirical evidence to inform policy, practice and investment.
The WHO global landscape of cancer clinical trials
Nature Medicine, Published online: 09 September 2025; doi:10.1038/s41591-025-03926-x
This Review of the WHO’s International Clinical Trials Registry Platform presents a snapshot of the global cancer trial landscape and provides critical empirical evidence to inform policy, practice and investment.-
Nature Medicine
-
Building the world’s first truly global medical foundation model
Nature Medicine, Published online: 08 September 2025; doi:10.1038/s41591-025-03859-5Building the world’s first truly global medical foundation model
Building the world’s first truly global medical foundation model
Nature Medicine, Published online: 08 September 2025; doi:10.1038/s41591-025-03859-5
Building the world’s first truly global medical foundation model-
InfoQ

-
Hugging Face Introduces AI Sheets, a No-Code Tool for Dataset Transformation
Hugging Face has released AI Sheets, an open-source application designed to let users build, transform, and enrich datasets using AI models through a spreadsheet-like interface. The tool, available both on the Hub and for local deployment, allows users to experiment with thousands of open models, including OpenAI’s gpt-oss, without requiring code. By Robert Krzaczyński
Hugging Face Introduces AI Sheets, a No-Code Tool for Dataset Transformation
Hugging Face has released AI Sheets, an open-source application designed to let users build, transform, and enrich datasets using AI models through a spreadsheet-like interface. The tool, available both on the Hub and for local deployment, allows users to experiment with thousands of open models, including OpenAI’s gpt-oss, without requiring code.
By Robert Krzaczyński-
Nature Biotechnology - Issue - nature.com science feeds
-
Antibody–bottlebrush prodrug conjugates for targeted cancer therapy
Nature Biotechnology, Published online: 09 September 2025; doi:10.1038/s41587-025-02772-zAntibody–bottlebrush conjugates expand the options for drug cargos compared to antibody–drug conjugates.
Antibody–bottlebrush prodrug conjugates for targeted cancer therapy
Nature Biotechnology, Published online: 09 September 2025; doi:10.1038/s41587-025-02772-z
Antibody–bottlebrush conjugates expand the options for drug cargos compared to antibody–drug conjugates.-
Omics In Lung
-
DNA methylation subtypes dictate metastatic heterogeneity of osteosarcoma via distinct tumor-stromal interactions: Multi-omics profiling and decitabine validation
Int J Biol Macromol. 2025 Sep 5;327(Pt 2):147473. doi: 10.1016/j.ijbiomac.2025.147473. Online ahead of print.ABSTRACTOsteosarcoma (OS), the most prevalent primary bone malignancy in adolescents, is characterized by aggressive progression and early metastasis. However, the epigenetic drivers of its metastatic heterogeneity remain poorly understood. Herein, we integrated bulk DNA methylation profiling and single-cell RNA sequencing (scRNA-seq) to elucidate the epigenetic mechanisms driving OS meta
DNA methylation subtypes dictate metastatic heterogeneity of osteosarcoma via distinct tumor-stromal interactions: Multi-omics profiling and decitabine validation
Int J Biol Macromol. 2025 Sep 5;327(Pt 2):147473. doi: 10.1016/j.ijbiomac.2025.147473. Online ahead of print.
ABSTRACT
Osteosarcoma (OS), the most prevalent primary bone malignancy in adolescents, is characterized by aggressive progression and early metastasis. However, the epigenetic drivers of its metastatic heterogeneity remain poorly understood. Herein, we integrated bulk DNA methylation profiling and single-cell RNA sequencing (scRNA-seq) to elucidate the epigenetic mechanisms driving OS metastatic heterogeneity. Consensus clustering identified two methylation subtypes (K = 2) with distinct survival outcomes, where hypermethylated (MSO-high) tumors exhibited poor prognosis. Weighted gene co-expression network analysis (WGCNA) revealed methylation-associated modules enriched in metabolic and immune pathways, pinpointing key genes such as CAMK1G and SLC11A1. Single-cell profiling uncovered MSO-high myeloid cells associated with inflammatory and oxidative phosphorylation pathways, while MSO-high OS cells displayed transdifferentiation toward fibroblasts via pseudotime trajectories, remodeling the extracellular matrix (ECM) to facilitate lung metastasis. Conversely, MSO-low tumors activated HLA-B-mediated neutrophil-CD8+ T cell interactions, promoting lymphatic metastasis via CXCR4/CXCL12 signaling. Furthermore, functional validation using the DNA demethylating agent decitabine demonstrated reduced fibroblastic transdifferentiation and suppressed invasive capacity in MSO-high osteosarcoma cells, supporting the therapeutic potential of targeting methylation dysregulation. These findings establish a model where DNA methylation dictates metastatic phenotypes through differential tumor-stromal crosstalk, providing novel targets for epigenetic therapy to disrupt fibrotic-immune networks and metastatic colonization.
PMID:40915448 | DOI:10.1016/j.ijbiomac.2025.147473
-
Omics In Lung
-
GASPS: A Multi-Omics Framework for Defining Genomic Aberration-Driven Signatures and Predicting Patient Outcomes in Lung Cancer
bioRxiv [Preprint]. 2025 Aug 25:2025.08.21.671519. doi: 10.1101/2025.08.21.671519.ABSTRACTLung cancer is the most common cause of cancer-related death worldwide. Recent advancements in targeted therapies and immunotherapies have achieved remarkable success. However, patient responses to treatments with lung cancer vary substantially. The mutation status of driver genes can direct personalized treatment, but their prognostic value and treatment efficacy are limited. In this study, we developed a
GASPS: A Multi-Omics Framework for Defining Genomic Aberration-Driven Signatures and Predicting Patient Outcomes in Lung Cancer
bioRxiv [Preprint]. 2025 Aug 25:2025.08.21.671519. doi: 10.1101/2025.08.21.671519.
ABSTRACT
Lung cancer is the most common cause of cancer-related death worldwide. Recent advancements in targeted therapies and immunotherapies have achieved remarkable success. However, patient responses to treatments with lung cancer vary substantially. The mutation status of driver genes can direct personalized treatment, but their prognostic value and treatment efficacy are limited. In this study, we developed a statistical framework named Genomic Aberration-Derived Signature for Patient Stratification (GASPS) to characterize the transcriptomic deregulation of driver genomic aberrations and stratify patients. By applying GASPS to The Cancer Genome Atlas Lung Adenocarcinoma (TCGA-LUAD) data, we developed gene signatures for 38 driver genomic aberrations, including gene mutations, amplifications, and deletions. These signatures were applied to independent lung cancer transcriptomic datasets containing a total of 2,226 patient samples. Our results indicated that these driver gene signatures are much more prognostic than their corresponding genomic mutations. Interestingly, the two EGFR-related signatures characterizing EGFR mutation and amplification, respectively, exhibited contrasting associations with prognosis, treatment response, and immune infiltration in the tumor microenvironment. Moreover, the STK11 mutation signature, rather than the mutation status, was found to be predictive of the response and long-term benefit of patients treated with immune checkpoint blockade therapy in lung cancer. This framework is readily applicable to most cancer types using existing data to improve prognostic risk assessment and treatment efficacy by guiding personalized therapies.
PMID:40909579 | PMC:PMC12407784 | DOI:10.1101/2025.08.21.671519
-
(Multiomics OR Omics) AND (Lung OR gastric OR Hepatocellular)
-
GASPS: A Multi-Omics Framework for Defining Genomic Aberration-Driven Signatures and Predicting Patient Outcomes in Lung Cancer
bioRxiv [Preprint]. 2025 Aug 25:2025.08.21.671519. doi: 10.1101/2025.08.21.671519.ABSTRACTLung cancer is the most common cause of cancer-related death worldwide. Recent advancements in targeted therapies and immunotherapies have achieved remarkable success. However, patient responses to treatments with lung cancer vary substantially. The mutation status of driver genes can direct personalized treatment, but their prognostic value and treatment efficacy are limited. In this study, we developed a
GASPS: A Multi-Omics Framework for Defining Genomic Aberration-Driven Signatures and Predicting Patient Outcomes in Lung Cancer
bioRxiv [Preprint]. 2025 Aug 25:2025.08.21.671519. doi: 10.1101/2025.08.21.671519.
ABSTRACT
Lung cancer is the most common cause of cancer-related death worldwide. Recent advancements in targeted therapies and immunotherapies have achieved remarkable success. However, patient responses to treatments with lung cancer vary substantially. The mutation status of driver genes can direct personalized treatment, but their prognostic value and treatment efficacy are limited. In this study, we developed a statistical framework named Genomic Aberration-Derived Signature for Patient Stratification (GASPS) to characterize the transcriptomic deregulation of driver genomic aberrations and stratify patients. By applying GASPS to The Cancer Genome Atlas Lung Adenocarcinoma (TCGA-LUAD) data, we developed gene signatures for 38 driver genomic aberrations, including gene mutations, amplifications, and deletions. These signatures were applied to independent lung cancer transcriptomic datasets containing a total of 2,226 patient samples. Our results indicated that these driver gene signatures are much more prognostic than their corresponding genomic mutations. Interestingly, the two EGFR-related signatures characterizing EGFR mutation and amplification, respectively, exhibited contrasting associations with prognosis, treatment response, and immune infiltration in the tumor microenvironment. Moreover, the STK11 mutation signature, rather than the mutation status, was found to be predictive of the response and long-term benefit of patients treated with immune checkpoint blockade therapy in lung cancer. This framework is readily applicable to most cancer types using existing data to improve prognostic risk assessment and treatment efficacy by guiding personalized therapies.
PMID:40909579 | PMC:PMC12407784 | DOI:10.1101/2025.08.21.671519
-
Cell
-
Scalable generation and functional classification of genetic variants in inborn errors of immunity to accelerate clinical diagnosis and treatment
In lieu of traditional genetic variant testing approaches, an approach using scalable variant classification in primary human T cells with a clinically relevant readout can inform rapid diagnosis and treatment of inborn errors of immunity.
Scalable generation and functional classification of genetic variants in inborn errors of immunity to accelerate clinical diagnosis and treatment
-
Cell
-
STAMP: Single-cell transcriptomics analysis and multimodal profiling through imaging
Single-cell transcriptomics analysis and multimodal profiling (STAMP) by imaging enables single-cell analysis of cells in suspension without the need for sequencing. The markedly reduced costs and flexible experimental designs support the profiling of millions of cells or the large-scale multiplexing of conditions, perturbations, and sample types.
STAMP: Single-cell transcriptomics analysis and multimodal profiling through imaging
-
Omics In Lung
-
Challenges in diagnosis of sarcoidosis
Curr Opin Immunol. 2025 Sep 1;97:102652. doi: 10.1016/j.coi.2025.102652. Online ahead of print.ABSTRACTPURPOSE OF REVIEW: Diagnosing sarcoidosis remains challenging. Histology findings and a variable clinical presentation can mimic other infectious, malignant, and autoimmune diseases. This review synthesizes current evidence on histopathology, sampling techniques, imaging modalities, and biomarkers and explores how emerging 'omics' and artificial intelligence tools may sharpen diagnostic accurac
Challenges in diagnosis of sarcoidosis
Curr Opin Immunol. 2025 Sep 1;97:102652. doi: 10.1016/j.coi.2025.102652. Online ahead of print.
ABSTRACT
PURPOSE OF REVIEW: Diagnosing sarcoidosis remains challenging. Histology findings and a variable clinical presentation can mimic other infectious, malignant, and autoimmune diseases. This review synthesizes current evidence on histopathology, sampling techniques, imaging modalities, and biomarkers and explores how emerging 'omics' and artificial intelligence tools may sharpen diagnostic accuracy.
RECENT FINDINGS: Within the typical granulomatous lesions, limited or 'burned-out' necrosis is an ancillary finding, which can be present in up to one-third of sarcoid biopsies, and demands a careful differential diagnostic work-up. Endobronchial ultrasound-guided transbronchial needle aspiration of lymph nodes has replaced mediastinoscopy as first-line sampling tool, while cryobiopsy is still under validation. Volumetric PET metrics such as total lung glycolysis and somatostatin-receptor tracers refine activity assessment; combined FDG PET/MRI improves detection of occult cardiac disease. Advanced bronchoalveolar lavage (BAL) immunophenotyping via flow cytometry and serum, BAL, and genetic biomarkers show to correlate with inflammatory burden but have low diagnostic value. Multi-omics signatures and Positron Emission Tomography with Computer Tomography radiomics, supported by deep-learning algorithms, show promising results for noninvasive diagnostic confirmation, phenotyping, and disease monitoring.
SUMMARY: No single test is conclusive for diagnosing sarcoidosis. An integrated, multidisciplinary strategy is needed. Large, multicenter, and multiethnic studies are essential to translate and validate data from emerging AI tools and -omics research into clinical routine.
PMID:40902264 | DOI:10.1016/j.coi.2025.102652
-
Journal of Medical Internet Research
-
Enabling Digital Compassion in Digital Health Environments: Modified eDelphi Study to Identify Interprofessional Competencies and Technology Attributes
Background: Health care continues to advance through digital innovation, and technology-enabled processes and interventions are increasingly being introduced to deliver and expand access to care. In this evolving digital health ecosystem, health care professionals (HCPs), learners, and organizations may not be prepared or equipped with the knowledge, skills, and behaviors required to navigate these new digital tools while simultaneously sustaining and integrating compassionate care. Moreover, th
Enabling Digital Compassion in Digital Health Environments: Modified eDelphi Study to Identify Interprofessional Competencies and Technology Attributes
-
npj Digital Medicine
-
Deep hierarchical subtyping of multi-organ systemic sclerosis trajectories - a EUSTAR study
npj Digital Medicine, Published online: 01 September 2025; doi:10.1038/s41746-025-01962-yDeep hierarchical subtyping of multi-organ systemic sclerosis trajectories - a EUSTAR study
Deep hierarchical subtyping of multi-organ systemic sclerosis trajectories - a EUSTAR study
npj Digital Medicine, Published online: 01 September 2025; doi:10.1038/s41746-025-01962-y
Deep hierarchical subtyping of multi-organ systemic sclerosis trajectories - a EUSTAR study-
(Multiomics OR Omics) AND (Lung OR gastric OR Hepatocellular)
-
Longitudinal liquid biopsy identifies an early predictive biomarker of immune checkpoint blockade response in head and neck squamous cell carcinoma
Nat Commun. 2025 Sep 1;16(1):8161. doi: 10.1038/s41467-025-63538-4.ABSTRACTImmune checkpoint blockade (ICB) has improved outcomes for patients with head and neck squamous cell carcinoma (HNSCC), but predictive biomarkers remain limited. Here, we use a time-resolved, multi-omic approach in a murine HNSCC model to characterize peripheral immune responses to ICB. Single-cell transcriptomics and T/B cell receptor analyses reveal early on-treatment expansion of effector memory T and B cell repertoire
Longitudinal liquid biopsy identifies an early predictive biomarker of immune checkpoint blockade response in head and neck squamous cell carcinoma
Nat Commun. 2025 Sep 1;16(1):8161. doi: 10.1038/s41467-025-63538-4.
ABSTRACT
Immune checkpoint blockade (ICB) has improved outcomes for patients with head and neck squamous cell carcinoma (HNSCC), but predictive biomarkers remain limited. Here, we use a time-resolved, multi-omic approach in a murine HNSCC model to characterize peripheral immune responses to ICB. Single-cell transcriptomics and T/B cell receptor analyses reveal early on-treatment expansion of effector memory T and B cell repertoires in responders, preceding tumor regression. These dynamic immune features inform a composite transcriptional signature that accurately predicts ICB response in independent human HNSCC cohorts. LiBIO outperforms existing biomarkers and generalizes to melanoma, non-small cell lung cancer, and breast cancer without retraining. These findings suggest that early treatment-induced changes in circulating immune repertoires reflect the host's capacity to mount an effective antitumor response. This work provides a framework for leveraging transient peripheral immune dynamics to develop non-invasive, high-fidelity biomarkers for response to immunotherapy across cancer types.
PMID:40890155 | PMC:PMC12402333 | DOI:10.1038/s41467-025-63538-4